Jump to
S1C2
Energy calculated at MP2=FULL/6-31G
| | hartrees |
|---|
| Energy at 0K | -167.902968 |
| Energy at 298.15K | |
| HF Energy | -167.532086 |
| Nuclear repulsion energy | 58.581178 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3539 |
3362 |
231.50 |
38.18 |
0.22 |
0.37 |
| 2 |
Σ |
2094 |
1989 |
436.63 |
7.25 |
0.07 |
0.13 |
| 3 |
Σ |
1217 |
1156 |
69.37 |
23.73 |
0.31 |
0.48 |
| 4 |
Π |
465 |
442 |
5.89 |
0.01 |
0.75 |
0.86 |
| 4 |
Π |
465 |
442 |
5.89 |
0.01 |
0.75 |
0.86 |
| 5 |
Π |
434i |
412i |
105.90 |
3.77 |
0.75 |
0.86 |
| 5 |
Π |
434i |
412i |
105.90 |
3.77 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 3455.5 cm
-1
Scaled (by 0.9501) Zero Point Vibrational Energy (zpe) 3283.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
1.221 |
| N2 |
0.000 |
0.000 |
-0.021 |
| C3 |
0.000 |
0.000 |
-1.223 |
| H4 |
0.000 |
0.000 |
-2.287 |
Atom - Atom Distances (Å)
| |
O1 |
N2 |
C3 |
H4 |
| O1 | | 1.2416 | 2.4434 | 3.5077 |
N2 | 1.2416 | | 1.2018 | 2.2661 | C3 | 2.4434 | 1.2018 | | 1.0643 | H4 | 3.5077 | 2.2661 | 1.0643 | |
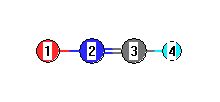 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
N2 |
C3 |
180.000 |
|
N2 |
C3 |
H4 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31G
| | hartrees |
|---|
| Energy at 0K | -167.904410 |
| Energy at 298.15K | -167.905188 |
| HF Energy | -167.524367 |
| Nuclear repulsion energy | 58.514985 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3480 |
3306 |
137.99 |
|
|
|
| 2 |
A' |
2095 |
1991 |
396.04 |
|
|
|
| 3 |
A' |
1229 |
1168 |
38.88 |
|
|
|
| 4 |
A' |
641 |
609 |
115.35 |
|
|
|
| 5 |
A' |
439 |
417 |
7.34 |
|
|
|
| 6 |
A" |
515 |
489 |
8.53 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4199.1 cm
-1
Scaled (by 0.9501) Zero Point Vibrational Energy (zpe) 3989.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-1.123 |
-0.464 |
0.000 |
| N2 |
0.000 |
0.055 |
0.000 |
| C3 |
1.174 |
0.368 |
0.000 |
| H4 |
1.940 |
1.114 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
N2 |
C3 |
H4 |
| O1 | | 1.2371 | 2.4432 | 3.4453 |
N2 | 1.2371 | | 1.2152 | 2.2100 | C3 | 2.4432 | 1.2152 | | 1.0690 | H4 | 3.4453 | 2.2100 | 1.0690 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
N2 |
C3 |
170.136 |
|
N2 |
C3 |
H4 |
150.652 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
