Jump to
S1C2
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -151.681237 |
| Energy at 298.15K | |
| HF Energy | -151.581363 |
| Nuclear repulsion energy | 51.593129 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3520 |
3520 |
101.55 |
49.28 |
0.22 |
0.36 |
| 2 |
Σ |
2048 |
2048 |
138.27 |
0.21 |
0.58 |
0.73 |
| 3 |
Σ |
1249 |
1249 |
35.87 |
25.97 |
0.26 |
0.42 |
| 4 |
Π |
494 |
494 |
1.99 |
1.08 |
0.75 |
0.86 |
| 4 |
Π |
463 |
463 |
0.54 |
1.14 |
0.75 |
0.86 |
| 5 |
Π |
422 |
422 |
35.91 |
0.63 |
0.75 |
0.86 |
| 5 |
Π |
299i |
299i |
160.62 |
6.43 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 3947.8 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 3947.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.008 |
| C2 |
0.000 |
0.000 |
-1.259 |
| O3 |
0.000 |
0.000 |
1.229 |
| H4 |
0.000 |
0.000 |
-2.322 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.2667 | 1.2211 | 2.3297 |
C2 | 1.2667 | | 2.4878 | 1.0630 | O3 | 1.2211 | 2.4878 | | 3.5508 | H4 | 2.3297 | 1.0630 | 3.5508 | |
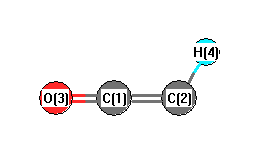 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
180.000 |
|
C2 |
C1 |
O3 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.123 |
|
|
|
| 2 |
C |
-0.057 |
|
|
|
| 3 |
O |
-0.333 |
|
|
|
| 4 |
H |
0.267 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.521 |
2.521 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.659 |
0.000 |
0.000 |
| y |
0.000 |
-16.028 |
0.000 |
| z |
0.000 |
0.000 |
-15.400 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.945 |
0.000 |
0.000 |
| y |
0.000 |
0.501 |
0.000 |
| z |
0.000 |
0.000 |
1.444 |
|
| Polar |
|---|
| 3z2-r2 | 2.887 |
|---|
| x2-y2 | -1.630 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.173 |
0.000 |
0.000 |
| y |
0.000 |
1.226 |
0.000 |
| z |
0.000 |
0.000 |
5.470 |
<r2> (average value of r
2) Å
2
| <r2> |
37.197 |
| (<r2>)1/2 |
6.099 |
Jump to
S1C1
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -151.681837 |
| Energy at 298.15K | |
| HF Energy | -151.579766 |
| Nuclear repulsion energy | 51.517142 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3470 |
3470 |
62.57 |
59.57 |
0.29 |
0.45 |
| 2 |
A' |
2035 |
2035 |
191.31 |
2.85 |
0.28 |
0.44 |
| 3 |
A' |
1239 |
1239 |
26.13 |
23.59 |
0.43 |
0.60 |
| 4 |
A' |
507 |
507 |
22.97 |
6.59 |
0.34 |
0.50 |
| 5 |
A' |
362 |
362 |
181.66 |
9.47 |
0.23 |
0.37 |
| 6 |
A" |
467 |
467 |
0.06 |
0.44 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 4040.2 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 4040.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.044 |
0.000 |
| C2 |
1.218 |
-0.358 |
0.000 |
| O3 |
-1.197 |
0.256 |
0.000 |
| H4 |
2.267 |
-0.158 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.2829 | 1.2158 | 2.2759 |
C2 | 1.2829 | | 2.4922 | 1.0674 | O3 | 1.2158 | 2.4922 | | 3.4887 | H4 | 2.2759 | 1.0674 | 3.4887 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
150.960 |
|
C2 |
C1 |
O3 |
171.790 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.231 |
|
|
|
| 2 |
C |
-0.117 |
|
|
|
| 3 |
O |
-0.328 |
|
|
|
| 4 |
H |
0.215 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.307 |
0.364 |
0.000 |
2.336 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-15.571 |
1.203 |
-0.005 |
| y |
1.203 |
-18.195 |
0.011 |
| z |
-0.005 |
0.011 |
-16.073 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.563 |
1.203 |
-0.005 |
| y |
1.203 |
-2.373 |
0.011 |
| z |
-0.005 |
0.011 |
0.810 |
|
| Polar |
|---|
| 3z2-r2 | 1.620 |
|---|
| x2-y2 | 2.624 |
|---|
| xy | 1.203 |
|---|
| xz | -0.005 |
|---|
| yz | 0.011 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.261 |
-1.114 |
0.001 |
| y |
-1.114 |
1.666 |
-0.001 |
| z |
0.001 |
-0.001 |
1.702 |
<r2> (average value of r
2) Å
2
| <r2> |
37.215 |
| (<r2>)1/2 |
6.100 |
