Jump to
S1C2
Energy calculated at MP2=FULL/6-31G
| | hartrees |
|---|
| Energy at 0K | -93.526623 |
| Energy at 298.15K | -93.527887 |
| HF Energy | -93.342312 |
| Nuclear repulsion energy | 28.420029 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3534 |
3357 |
62.58 |
|
|
|
| 2 |
A' |
2983 |
2834 |
709.94 |
|
|
|
| 3 |
A' |
2566 |
2438 |
358.92 |
|
|
|
| 4 |
A' |
1038 |
986 |
52.60 |
|
|
|
| 5 |
A' |
721 |
685 |
515.15 |
|
|
|
| 6 |
A" |
994 |
944 |
44.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5917.3 cm
-1
Scaled (by 0.9501) Zero Point Vibrational Energy (zpe) 5622.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.098 |
0.639 |
0.000 |
| N2 |
0.098 |
-0.564 |
0.000 |
| H3 |
-0.684 |
1.428 |
0.000 |
| H4 |
-0.588 |
-1.316 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.2036 | 1.1103 | 2.0725 |
N2 | 1.2036 | | 2.1399 | 1.0181 | H3 | 1.1103 | 2.1399 | | 2.7457 | H4 | 2.0725 | 1.0181 | 2.7457 | |
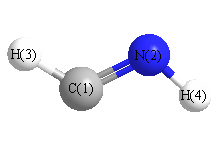 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
137.620 |
|
H3 |
C1 |
N2 |
135.241 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/6-31G
| | hartrees |
|---|
| Energy at 0K | -93.534952 |
| Energy at 298.15K | -93.536289 |
| HF Energy | -93.350264 |
| Nuclear repulsion energy | 28.353075 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3615 |
3435 |
97.55 |
|
|
|
| 2 |
A' |
3043 |
2892 |
415.04 |
|
|
|
| 3 |
A' |
2576 |
2448 |
597.40 |
|
|
|
| 4 |
A' |
1221 |
1160 |
13.75 |
|
|
|
| 5 |
A' |
838 |
796 |
603.30 |
|
|
|
| 6 |
A" |
1045 |
993 |
266.93 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6169.3 cm
-1
Scaled (by 0.9501) Zero Point Vibrational Energy (zpe) 5861.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.008 |
0.649 |
0.000 |
| N2 |
0.008 |
-0.566 |
0.000 |
| H3 |
-0.897 |
1.283 |
0.000 |
| H4 |
0.791 |
-1.213 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.2154 | 1.1047 | 2.0204 |
N2 | 1.2154 | | 2.0585 | 1.0159 | H3 | 1.1047 | 2.0585 | | 3.0132 | H4 | 2.0204 | 1.0159 | 3.0132 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
129.555 |
|
H3 |
C1 |
N2 |
124.990 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
