Jump to
S1C2
Energy calculated at M06-2X/6-31G
| | hartrees |
|---|
| Energy at 0K | -93.873132 |
| Energy at 298.15K | -93.874411 |
| HF Energy | -93.873132 |
| Nuclear repulsion energy | 28.080186 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3375 |
3375 |
0.89 |
|
|
|
| 2 |
A' |
3044 |
3044 |
44.22 |
|
|
|
| 3 |
A' |
1906 |
1906 |
35.29 |
|
|
|
| 4 |
A' |
1001 |
1001 |
63.69 |
|
|
|
| 5 |
A' |
807 |
807 |
290.60 |
|
|
|
| 6 |
A" |
949 |
949 |
17.60 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5540.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 5540.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.104 |
0.647 |
0.000 |
| N2 |
0.104 |
-0.580 |
0.000 |
| H3 |
-0.674 |
1.422 |
0.000 |
| H4 |
-0.677 |
-1.245 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.2279 | 1.0981 | 2.0467 |
N2 | 1.2279 | | 2.1486 | 1.0249 | H3 | 1.0981 | 2.1486 | | 2.6669 | H4 | 2.0467 | 1.0249 | 2.6669 | |
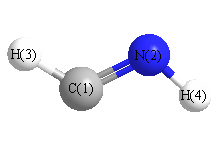 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
130.395 |
|
H3 |
C1 |
N2 |
134.881 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.019 |
|
|
|
| 2 |
N |
-0.490 |
|
|
|
| 3 |
H |
0.155 |
|
|
|
| 4 |
H |
0.317 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.547 |
0.132 |
0.000 |
2.550 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-12.428 |
0.801 |
0.000 |
| y |
0.801 |
-10.619 |
0.000 |
| z |
0.000 |
0.000 |
-12.492 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.873 |
0.801 |
0.000 |
| y |
0.801 |
1.842 |
0.000 |
| z |
0.000 |
0.000 |
-0.969 |
|
| Polar |
|---|
| 3z2-r2 | -1.938 |
|---|
| x2-y2 | -1.810 |
|---|
| xy | 0.801 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.682 |
0.065 |
0.000 |
| y |
0.065 |
4.055 |
0.000 |
| z |
0.000 |
0.000 |
1.059 |
<r2> (average value of r
2) Å
2
| <r2> |
16.897 |
| (<r2>)1/2 |
4.111 |
Jump to
S1C1
Energy calculated at M06-2X/6-31G
| | hartrees |
|---|
| Energy at 0K | -93.881199 |
| Energy at 298.15K | -93.882548 |
| HF Energy | -93.881199 |
| Nuclear repulsion energy | 27.994808 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3515 |
3515 |
8.95 |
|
|
|
| 2 |
A' |
3108 |
3108 |
23.41 |
|
|
|
| 3 |
A' |
1830 |
1830 |
28.76 |
|
|
|
| 4 |
A' |
1202 |
1202 |
9.80 |
|
|
|
| 5 |
A' |
922 |
922 |
278.28 |
|
|
|
| 6 |
A" |
1010 |
1010 |
155.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5793.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 5793.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.003 |
0.657 |
0.000 |
| N2 |
-0.003 |
-0.583 |
0.000 |
| H3 |
0.890 |
1.290 |
0.000 |
| H4 |
-0.850 |
-1.153 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.2404 | 1.0949 | 1.9990 |
N2 | 1.2404 | | 2.0755 | 1.0210 | H3 | 1.0949 | 2.0755 | | 3.0000 | H4 | 1.9990 | 1.0210 | 3.0000 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
123.955 |
|
H3 |
C1 |
N2 |
125.309 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.015 |
|
|
|
| 2 |
N |
-0.486 |
|
|
|
| 3 |
H |
0.170 |
|
|
|
| 4 |
H |
0.330 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.614 |
0.068 |
0.000 |
0.618 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-11.367 |
3.536 |
0.000 |
| y |
3.536 |
-11.207 |
0.000 |
| z |
0.000 |
0.000 |
-12.537 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.505 |
3.536 |
0.000 |
| y |
3.536 |
0.745 |
0.000 |
| z |
0.000 |
0.000 |
-1.250 |
|
| Polar |
|---|
| 3z2-r2 | -2.500 |
|---|
| x2-y2 | -0.160 |
|---|
| xy | 3.536 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.045 |
0.424 |
0.000 |
| y |
0.424 |
3.740 |
0.000 |
| z |
0.000 |
0.000 |
1.091 |
<r2> (average value of r
2) Å
2
| <r2> |
16.793 |
| (<r2>)1/2 |
4.098 |
