return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for HO2CCHNH2CH2OH (Serine)

using model chemistry: MP2=FULL/6-31G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-31G
 hartrees
Energy at 0K-397.291224
Energy at 298.15K-397.301141
HF Energy-396.525898
Nuclear repulsion energy319.587315
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3662 3479 6.50      
2 A 3636 3455 25.59      
3 A 3592 3412 46.64      
4 A 3536 3359 2.54      
5 A 3187 3028 15.57      
6 A 3103 2949 8.58      
7 A 3048 2896 41.19      
8 A 1748 1661 41.65      
9 A 1683 1599 133.39      
10 A 1573 1494 2.95      
11 A 1438 1366 10.57      
12 A 1420 1349 4.00      
13 A 1414 1343 18.69      
14 A 1394 1325 31.59      
15 A 1341 1274 22.96      
16 A 1269 1205 9.52      
17 A 1191 1131 14.16      
18 A 1160 1103 24.65      
19 A 1120 1064 216.03      
20 A 1046 994 46.73      
21 A 1010 960 57.36      
22 A 990 940 67.69      
23 A 810 769 57.71      
24 A 750 712 245.30      
25 A 713 678 29.60      
26 A 628 597 163.61      
27 A 585 556 9.87      
28 A 524 498 296.92      
29 A 505 480 14.60      
30 A 446 423 5.70      
31 A 302 287 9.57      
32 A 283 269 36.86      
33 A 275 261 17.93      
34 A 212 202 1.67      
35 A 165 157 4.64      
36 A 41 39 1.46      

Unscaled Zero Point Vibrational Energy (zpe) 24898.8 cm-1
Scaled (by 0.9501) Zero Point Vibrational Energy (zpe) 23656.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G
ABC
0.11374 0.07565 0.05020

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.781 -0.560 0.013
O2 2.097 -0.379 -0.372
O3 0.375 -1.598 0.567
C4 -0.075 0.678 -0.307
C5 -1.520 0.478 0.194
O6 -2.170 -0.674 -0.397
N7 0.482 1.942 0.228
H8 2.664 -1.157 -0.176
H9 -0.112 0.761 -1.402
H10 -1.514 0.427 1.298
H11 -2.091 1.365 -0.095
H12 -1.799 -1.510 -0.056
H13 1.369 2.207 -0.199
H14 0.579 1.944 1.244

Atom - Atom Distances (Å)
  C1 O2 O3 C4 C5 O6 N7 H8 H9 H10 H11 H12 H13 H14
C11.38311.24501.53872.53132.98122.52911.98442.13192.80923.45912.75022.83712.7974
O21.38312.30972.41623.76054.27692.89060.98232.69084.05934.54524.06912.69243.2114
O31.24502.30972.47962.83622.87393.55832.44703.11162.86423.91142.26314.00733.6118
C41.53872.41622.47961.54312.49501.48103.29921.09872.16992.14072.79742.10592.1058
C52.53133.76052.83621.54311.44862.48074.50782.14721.10451.09302.02373.39032.7671
O62.98124.27692.87392.49501.44863.77694.86292.70252.12432.06210.97644.56774.1351
N72.52912.89063.55831.48102.48073.77693.81132.09802.72482.65714.14771.01951.0213
H81.98440.98232.44703.29924.50784.86293.81133.59014.70495.38304.47863.60463.9970
H92.13192.69083.11161.09872.14722.70252.09803.59013.05982.44723.13312.39402.9793
H102.80924.05932.86422.16991.10452.12432.72484.70493.05981.77502.38073.70432.5851
H113.45914.54523.91142.14071.09302.06212.65715.38302.44721.77502.89003.56323.0428
H122.75024.06912.26312.79742.02370.97644.14774.47863.13312.38072.89004.88664.3903
H132.83712.69244.00732.10593.39034.56771.01953.60462.39403.70433.56324.88661.6666
H142.79743.21143.61182.10582.76714.13511.02133.99702.97932.58513.04284.39031.6666

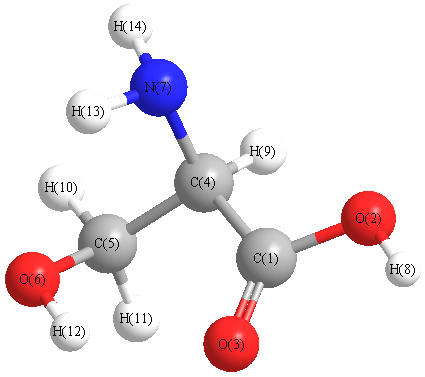
picture of Serine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 O2 H8 112.960 C1 C4 C5 110.449
C1 C4 N7 113.751 C1 C4 H9 106.676
O2 C1 O3 122.917 O2 C1 C4 111.466
O3 C1 C4 125.607 C4 C5 O6 112.985
C4 C5 H10 108.957 C4 C5 H11 107.369
C4 N7 H13 113.464 C4 N7 H14 113.331
C5 C4 N7 110.221 C5 C4 H9 107.544
C5 O6 H12 111.649 O6 C5 H10 111.918
O6 C5 H11 107.628 N7 C4 H9 107.921
H10 C5 H11 107.745 H13 N7 H14 109.498
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability