Jump to
S1C2
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -190.348158 |
| Energy at 298.15K | |
| HF Energy | -190.207744 |
| Nuclear repulsion energy | 87.196137 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3117 |
3117 |
29.70 |
204.67 |
0.11 |
0.20 |
| 2 |
A1 |
2283 |
2283 |
620.29 |
24.68 |
0.34 |
0.51 |
| 3 |
A1 |
1727 |
1727 |
35.12 |
4.51 |
0.48 |
0.65 |
| 4 |
A1 |
1531 |
1531 |
9.35 |
22.29 |
0.51 |
0.68 |
| 5 |
A1 |
904 |
904 |
8.73 |
42.57 |
0.29 |
0.45 |
| 6 |
B1 |
1046 |
1046 |
28.74 |
0.18 |
0.75 |
0.86 |
| 7 |
B1 |
588 |
588 |
17.15 |
3.76 |
0.75 |
0.86 |
| 8 |
B1 |
222 |
222 |
0.05 |
4.32 |
0.75 |
0.86 |
| 9 |
B2 |
3190 |
3190 |
17.39 |
132.08 |
0.75 |
0.86 |
| 10 |
B2 |
1108 |
1108 |
0.90 |
0.10 |
0.75 |
0.86 |
| 11 |
B2 |
359 |
359 |
7.68 |
1.02 |
0.75 |
0.86 |
| 12 |
B2 |
156i |
156i |
19.12 |
0.41 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 7958.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 7958.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.893 |
| C2 |
0.000 |
0.000 |
-0.566 |
| C3 |
0.000 |
0.000 |
0.715 |
| O4 |
0.000 |
0.000 |
1.928 |
| H5 |
0.000 |
0.921 |
-2.478 |
| H6 |
0.000 |
-0.921 |
-2.478 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3273 | 2.6082 | 3.8212 | 1.0908 | 1.0908 |
C2 | 1.3273 | | 1.2808 | 2.4939 | 2.1220 | 2.1220 | C3 | 2.6082 | 1.2808 | | 1.2130 | 3.3227 | 3.3227 | O4 | 3.8212 | 2.4939 | 1.2130 | | 4.5008 | 4.5008 | H5 | 1.0908 | 2.1220 | 3.3227 | 4.5008 | | 1.8420 | H6 | 1.0908 | 2.1220 | 3.3227 | 4.5008 | 1.8420 | |
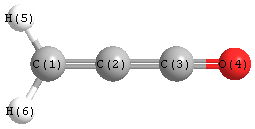 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
180.000 |
|
C2 |
C1 |
H5 |
122.394 |
| C2 |
C1 |
H6 |
122.394 |
|
C2 |
C3 |
O4 |
180.000 |
| H5 |
C1 |
H6 |
115.212 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.258 |
|
|
|
| 2 |
C |
0.169 |
|
|
|
| 3 |
C |
0.066 |
|
|
|
| 4 |
O |
-0.319 |
|
|
|
| 5 |
H |
0.171 |
|
|
|
| 6 |
H |
0.171 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-3.110 |
3.110 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.480 |
0.000 |
0.000 |
| y |
0.000 |
-22.431 |
0.000 |
| z |
0.000 |
0.000 |
-22.893 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.182 |
0.000 |
0.000 |
| y |
0.000 |
0.256 |
0.000 |
| z |
0.000 |
0.000 |
-0.438 |
|
| Polar |
|---|
| 3z2-r2 | -0.876 |
|---|
| x2-y2 | -0.049 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.589 |
0.000 |
0.000 |
| y |
0.000 |
2.489 |
0.000 |
| z |
0.000 |
0.000 |
10.594 |
<r2> (average value of r
2) Å
2
| <r2> |
84.310 |
| (<r2>)1/2 |
9.182 |
Jump to
S1C1
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -190.348606 |
| Energy at 298.15K | |
| HF Energy | -190.206780 |
| Nuclear repulsion energy | 87.229912 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3201 |
3201 |
16.07 |
131.83 |
0.68 |
0.81 |
| 2 |
A' |
3121 |
3121 |
27.35 |
180.35 |
0.12 |
0.21 |
| 3 |
A' |
2221 |
2221 |
615.48 |
29.73 |
0.37 |
0.54 |
| 4 |
A' |
1724 |
1724 |
29.34 |
3.57 |
0.55 |
0.71 |
| 5 |
A' |
1531 |
1531 |
7.08 |
22.67 |
0.53 |
0.69 |
| 6 |
A' |
1100 |
1100 |
7.19 |
2.44 |
0.21 |
0.34 |
| 7 |
A' |
914 |
914 |
5.92 |
39.32 |
0.25 |
0.40 |
| 8 |
A' |
427 |
427 |
12.64 |
3.51 |
0.56 |
0.71 |
| 9 |
A' |
152 |
152 |
17.11 |
3.69 |
0.70 |
0.83 |
| 10 |
A" |
1037 |
1037 |
28.82 |
0.30 |
0.75 |
0.86 |
| 11 |
A" |
644 |
644 |
14.55 |
4.71 |
0.75 |
0.86 |
| 12 |
A" |
246 |
246 |
0.52 |
4.99 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 8158.2 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 8158.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.696 |
-1.740 |
0.000 |
| C2 |
0.000 |
-0.605 |
0.000 |
| C3 |
-0.220 |
0.670 |
0.000 |
| O4 |
-0.607 |
1.817 |
0.000 |
| H5 |
1.787 |
-1.769 |
0.000 |
| H6 |
0.205 |
-2.712 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
| C1 | | 1.3319 | 2.5779 | 3.7879 | 1.0916 | 1.0887 |
C2 | 1.3319 | | 1.2930 | 2.4961 | 2.1329 | 2.1172 | C3 | 2.5779 | 1.2930 | | 1.2106 | 3.1578 | 3.4081 | O4 | 3.7879 | 2.4961 | 1.2106 | | 4.3109 | 4.6007 | H5 | 1.0916 | 2.1329 | 3.1578 | 4.3109 | | 1.8421 | H6 | 1.0887 | 2.1172 | 3.4081 | 4.6007 | 1.8421 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
158.262 |
|
C2 |
C1 |
H5 |
122.999 |
| C2 |
C1 |
H6 |
121.683 |
|
C2 |
C3 |
O4 |
171.132 |
| H5 |
C1 |
H6 |
115.318 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.280 |
|
|
|
| 2 |
C |
0.089 |
|
|
|
| 3 |
C |
0.173 |
|
|
|
| 4 |
O |
-0.321 |
|
|
|
| 5 |
H |
0.172 |
|
|
|
| 6 |
H |
0.167 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.489 |
-2.602 |
0.000 |
2.998 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.255 |
-0.454 |
-0.001 |
| y |
-0.454 |
-23.151 |
0.003 |
| z |
-0.001 |
0.003 |
-22.529 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.584 |
-0.454 |
-0.001 |
| y |
-0.454 |
-0.758 |
0.003 |
| z |
-0.001 |
0.003 |
0.174 |
|
| Polar |
|---|
| 3z2-r2 | 0.348 |
|---|
| x2-y2 | 0.895 |
|---|
| xy | -0.454 |
|---|
| xz | -0.001 |
|---|
| yz | 0.003 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.545 |
-2.374 |
0.000 |
| y |
-2.374 |
9.599 |
-0.000 |
| z |
0.000 |
-0.000 |
1.622 |
<r2> (average value of r
2) Å
2
| <r2> |
83.457 |
| (<r2>)1/2 |
9.135 |
