Jump to
S1C2
Vibrational Frequencies calculated at B2PLYP/6-31G
Geometric Data calculated at B2PLYP/6-31G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -553.671268 |
| Energy at 298.15K | -553.676646 |
| HF Energy | -553.528277 |
| Nuclear repulsion energy | 197.663591 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3282 |
3282 |
11.25 |
|
|
|
| 2 |
A |
3281 |
3281 |
9.78 |
|
|
|
| 3 |
A |
3241 |
3241 |
5.75 |
|
|
|
| 4 |
A |
3226 |
3226 |
6.04 |
|
|
|
| 5 |
A |
3189 |
3189 |
1.85 |
|
|
|
| 6 |
A |
3186 |
3186 |
2.60 |
|
|
|
| 7 |
A |
1675 |
1675 |
27.48 |
|
|
|
| 8 |
A |
1667 |
1667 |
25.10 |
|
|
|
| 9 |
A |
1472 |
1472 |
4.08 |
|
|
|
| 10 |
A |
1468 |
1468 |
8.95 |
|
|
|
| 11 |
A |
1340 |
1340 |
3.91 |
|
|
|
| 12 |
A |
1331 |
1331 |
20.09 |
|
|
|
| 13 |
A |
1089 |
1089 |
12.92 |
|
|
|
| 14 |
A |
1069 |
1069 |
9.53 |
|
|
|
| 15 |
A |
1022 |
1022 |
26.41 |
|
|
|
| 16 |
A |
1011 |
1011 |
54.27 |
|
|
|
| 17 |
A |
980 |
980 |
31.36 |
|
|
|
| 18 |
A |
953 |
953 |
44.53 |
|
|
|
| 19 |
A |
694 |
694 |
14.73 |
|
|
|
| 20 |
A |
653 |
653 |
0.92 |
|
|
|
| 21 |
A |
632 |
632 |
19.83 |
|
|
|
| 22 |
A |
613 |
613 |
13.63 |
|
|
|
| 23 |
A |
448 |
448 |
0.52 |
|
|
|
| 24 |
A |
371 |
371 |
0.73 |
|
|
|
| 25 |
A |
206 |
206 |
1.05 |
|
|
|
| 26 |
A |
130 |
130 |
3.37 |
|
|
|
| 27 |
A |
78 |
78 |
1.72 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19153.2 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 19153.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.006 |
-1.027 |
-0.011 |
| C2 |
1.127 |
0.368 |
0.404 |
| C3 |
-1.619 |
-0.168 |
-0.015 |
| C4 |
2.287 |
0.581 |
-0.229 |
| C5 |
-1.824 |
1.151 |
-0.131 |
| H6 |
0.797 |
0.963 |
1.250 |
| H7 |
-2.431 |
-0.877 |
0.094 |
| H8 |
2.964 |
1.361 |
0.104 |
| H9 |
2.599 |
-0.014 |
-1.081 |
| H10 |
-1.016 |
1.862 |
-0.266 |
| H11 |
-2.832 |
1.552 |
-0.102 |
Atom - Atom Distances (Å)
| |
S1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| S1 | | 1.8372 | 1.8376 | 2.7998 | 2.8474 | 2.4849 | 2.4439 | 3.8042 | 2.9827 | 3.0750 | 3.8358 |
C2 | 1.8372 | | 2.8284 | 1.3390 | 3.0997 | 1.0855 | 3.7823 | 2.1104 | 2.1257 | 2.6968 | 4.1631 | C3 | 1.8376 | 2.8284 | | 3.9824 | 1.3398 | 2.9516 | 1.0843 | 4.8326 | 4.3526 | 2.1320 | 2.1066 | C4 | 2.7998 | 1.3390 | 3.9824 | | 4.1516 | 2.1344 | 4.9491 | 1.0857 | 1.0847 | 3.5428 | 5.2119 | C5 | 2.8474 | 3.0997 | 1.3398 | 4.1516 | | 2.9684 | 2.1292 | 4.7986 | 4.6712 | 1.0846 | 1.0852 | H6 | 2.4849 | 1.0855 | 2.9516 | 2.1344 | 2.9684 | | 3.8909 | 2.4842 | 3.1042 | 2.5282 | 3.9171 | H7 | 2.4439 | 3.7823 | 1.0843 | 4.9491 | 2.1292 | 3.8909 | | 5.8414 | 5.2372 | 3.1041 | 2.4698 | H8 | 3.8042 | 2.1104 | 4.8326 | 1.0857 | 4.7986 | 2.4842 | 5.8414 | | 1.8518 | 4.0284 | 5.8030 | H9 | 2.9827 | 2.1257 | 4.3526 | 1.0847 | 4.6712 | 3.1042 | 5.2372 | 1.8518 | | 4.1531 | 5.7362 | H10 | 3.0750 | 2.6968 | 2.1320 | 3.5428 | 1.0846 | 2.5282 | 3.1041 | 4.0284 | 4.1531 | | 1.8497 | H11 | 3.8358 | 4.1631 | 2.1066 | 5.2119 | 1.0852 | 3.9171 | 2.4698 | 5.8030 | 5.7362 | 1.8497 | |
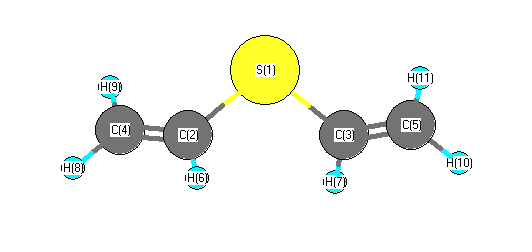 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
C2 |
C4 |
122.869 |
|
S1 |
C2 |
H6 |
113.986 |
| S1 |
C3 |
C5 |
126.598 |
|
S1 |
C3 |
H7 |
110.872 |
| C2 |
S1 |
C3 |
100.649 |
|
C2 |
C4 |
H8 |
120.640 |
| C2 |
C4 |
H9 |
122.228 |
|
C3 |
C5 |
H10 |
122.789 |
| C3 |
C5 |
H11 |
120.254 |
|
C4 |
C2 |
H6 |
123.029 |
| C5 |
C3 |
H7 |
122.530 |
|
H8 |
C4 |
H9 |
117.129 |
| H10 |
C5 |
H11 |
116.955 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
0.321 |
|
|
|
| 2 |
C |
-0.366 |
|
|
|
| 3 |
C |
-0.366 |
|
|
|
| 4 |
C |
-0.259 |
|
|
|
| 5 |
C |
-0.276 |
|
|
|
| 6 |
H |
0.173 |
|
|
|
| 7 |
H |
0.171 |
|
|
|
| 8 |
H |
0.149 |
|
|
|
| 9 |
H |
0.157 |
|
|
|
| 10 |
H |
0.149 |
|
|
|
| 11 |
H |
0.147 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.085 |
1.456 |
0.258 |
1.481 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.653 |
1.691 |
-0.771 |
| y |
1.691 |
-38.362 |
1.031 |
| z |
-0.771 |
1.031 |
-40.442 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.749 |
1.691 |
-0.771 |
| y |
1.691 |
-1.815 |
1.031 |
| z |
-0.771 |
1.031 |
-4.935 |
|
| Polar |
|---|
| 3z2-r2 | -9.869 |
|---|
| x2-y2 | 5.709 |
|---|
| xy | 1.691 |
|---|
| xz | -0.771 |
|---|
| yz | 1.031 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.031 |
0.223 |
-0.858 |
| y |
0.223 |
8.780 |
0.280 |
| z |
-0.858 |
0.280 |
4.635 |
<r2> (average value of r
2) Å
2
| <r2> |
170.526 |
| (<r2>)1/2 |
13.059 |
