Jump to
S1C2
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -189.462092 |
| Energy at 298.15K | -189.464779 |
| HF Energy | -189.341954 |
| Nuclear repulsion energy | 68.556969 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3599 |
3599 |
34.27 |
90.43 |
0.30 |
0.47 |
| 2 |
A' |
3190 |
3190 |
33.86 |
110.03 |
0.27 |
0.43 |
| 3 |
A' |
1700 |
1700 |
244.97 |
4.74 |
0.25 |
0.40 |
| 4 |
A' |
1420 |
1420 |
5.84 |
11.80 |
0.62 |
0.76 |
| 5 |
A' |
1303 |
1303 |
8.49 |
1.21 |
0.64 |
0.78 |
| 6 |
A' |
1068 |
1068 |
264.11 |
4.45 |
0.16 |
0.27 |
| 7 |
A' |
601 |
601 |
47.93 |
6.31 |
0.63 |
0.78 |
| 8 |
A" |
1047 |
1047 |
7.63 |
1.62 |
0.75 |
0.86 |
| 9 |
A" |
683 |
683 |
218.43 |
3.47 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 7305.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 7305.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.435 |
0.000 |
| O2 |
-1.052 |
-0.454 |
0.000 |
| O3 |
1.192 |
0.119 |
0.000 |
| H4 |
-0.390 |
1.452 |
0.000 |
| H5 |
-0.732 |
-1.383 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
H4 |
H5 |
| C1 | | 1.3777 | 1.2335 | 1.0888 | 1.9606 |
O2 | 1.3777 | | 2.3163 | 2.0177 | 0.9829 | O3 | 1.2335 | 2.3163 | | 2.0689 | 2.4413 | H4 | 1.0888 | 2.0177 | 2.0689 | | 2.8559 | H5 | 1.9606 | 0.9829 | 2.4413 | 2.8559 | |
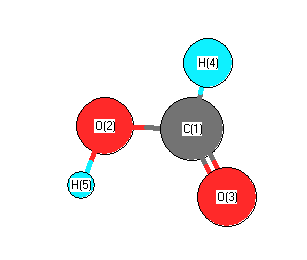 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
111.212 |
|
O2 |
C1 |
O3 |
124.923 |
| O2 |
C1 |
H4 |
109.218 |
|
O3 |
C1 |
H4 |
125.859 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.372 |
|
|
|
| 2 |
O |
-0.551 |
|
|
|
| 3 |
O |
-0.389 |
|
|
|
| 4 |
H |
0.175 |
|
|
|
| 5 |
H |
0.393 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.273 |
-0.429 |
0.000 |
1.343 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.234 |
0.085 |
-0.001 |
| y |
0.085 |
-12.369 |
-0.002 |
| z |
-0.001 |
-0.002 |
-16.955 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.572 |
0.085 |
-0.001 |
| y |
0.085 |
7.726 |
-0.002 |
| z |
-0.001 |
-0.002 |
0.846 |
|
| Polar |
|---|
| 3z2-r2 | 1.693 |
|---|
| x2-y2 | -10.865 |
|---|
| xy | 0.085 |
|---|
| xz | -0.001 |
|---|
| yz | -0.002 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.264 |
0.134 |
-0.000 |
| y |
0.134 |
2.719 |
0.000 |
| z |
-0.000 |
0.000 |
1.055 |
<r2> (average value of r
2) Å
2
| <r2> |
38.778 |
| (<r2>)1/2 |
6.227 |
Jump to
S1C1
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -189.452409 |
| Energy at 298.15K | |
| HF Energy | -189.332228 |
| Nuclear repulsion energy | 68.462439 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3653 |
3653 |
33.82 |
124.65 |
0.30 |
0.46 |
| 2 |
A' |
3063 |
3063 |
77.05 |
97.10 |
0.29 |
0.45 |
| 3 |
A' |
1763 |
1763 |
195.32 |
6.72 |
0.15 |
0.26 |
| 4 |
A' |
1450 |
1450 |
1.52 |
11.70 |
0.64 |
0.78 |
| 5 |
A' |
1210 |
1210 |
285.20 |
7.66 |
0.71 |
0.83 |
| 6 |
A' |
1083 |
1083 |
83.44 |
9.65 |
0.29 |
0.45 |
| 7 |
A' |
636 |
636 |
10.82 |
1.81 |
0.71 |
0.83 |
| 8 |
A" |
1027 |
1027 |
0.05 |
1.96 |
0.75 |
0.86 |
| 9 |
A" |
504 |
504 |
143.47 |
3.50 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 7193.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 7193.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.401 |
0.000 |
| O2 |
-0.921 |
-0.631 |
0.000 |
| O3 |
1.208 |
0.194 |
0.000 |
| H4 |
-0.449 |
1.403 |
0.000 |
| H5 |
-1.846 |
-0.316 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
H4 |
H5 |
| C1 | | 1.3827 | 1.2253 | 1.0980 | 1.9804 |
O2 | 1.3827 | | 2.2826 | 2.0873 | 0.9776 | O3 | 1.2253 | 2.2826 | | 2.0510 | 3.0961 | H4 | 1.0980 | 2.0873 | 2.0510 | | 2.2146 | H5 | 1.9804 | 0.9776 | 3.0961 | 2.2146 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
112.954 |
|
O2 |
C1 |
O3 |
122.031 |
| O2 |
C1 |
H4 |
114.096 |
|
O3 |
C1 |
H4 |
123.872 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.381 |
|
|
|
| 2 |
O |
-0.545 |
|
|
|
| 3 |
O |
-0.359 |
|
|
|
| 4 |
H |
0.134 |
|
|
|
| 5 |
H |
0.390 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.836 |
1.990 |
0.000 |
4.321 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-15.634 |
-1.287 |
-0.001 |
| y |
-1.287 |
-17.861 |
-0.001 |
| z |
-0.001 |
-0.001 |
-16.952 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.773 |
-1.287 |
-0.001 |
| y |
-1.287 |
-1.568 |
-0.001 |
| z |
-0.001 |
-0.001 |
-0.205 |
|
| Polar |
|---|
| 3z2-r2 | -0.410 |
|---|
| x2-y2 | 2.228 |
|---|
| xy | -1.287 |
|---|
| xz | -0.001 |
|---|
| yz | -0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.935 |
0.164 |
-0.000 |
| y |
0.164 |
2.221 |
0.000 |
| z |
-0.000 |
0.000 |
1.056 |
<r2> (average value of r
2) Å
2
| <r2> |
39.077 |
| (<r2>)1/2 |
6.251 |
