Jump to
S1C2
Vibrational Frequencies calculated at B2PLYP/6-31G
Geometric Data calculated at B2PLYP/6-31G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -193.995881 |
| Energy at 298.15K | |
| HF Energy | -193.862373 |
| Nuclear repulsion energy | 131.587634 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3664 |
3664 |
3.71 |
142.23 |
0.31 |
0.48 |
| 2 |
A |
3168 |
3168 |
23.91 |
37.54 |
0.73 |
0.84 |
| 3 |
A |
3133 |
3133 |
60.09 |
52.52 |
0.56 |
0.72 |
| 4 |
A |
3106 |
3106 |
23.79 |
75.11 |
0.75 |
0.86 |
| 5 |
A |
3068 |
3068 |
67.63 |
116.66 |
0.17 |
0.28 |
| 6 |
A |
3061 |
3061 |
51.79 |
119.54 |
0.11 |
0.20 |
| 7 |
A |
3054 |
3054 |
9.96 |
70.81 |
0.47 |
0.64 |
| 8 |
A |
3020 |
3020 |
51.75 |
90.07 |
0.17 |
0.28 |
| 9 |
A |
1581 |
1581 |
2.21 |
8.99 |
0.69 |
0.82 |
| 10 |
A |
1575 |
1575 |
8.31 |
11.66 |
0.75 |
0.86 |
| 11 |
A |
1558 |
1558 |
7.68 |
25.19 |
0.75 |
0.86 |
| 12 |
A |
1547 |
1547 |
2.20 |
20.71 |
0.75 |
0.86 |
| 13 |
A |
1482 |
1482 |
8.04 |
2.56 |
0.60 |
0.75 |
| 14 |
A |
1462 |
1462 |
2.93 |
3.68 |
0.62 |
0.76 |
| 15 |
A |
1435 |
1435 |
1.31 |
2.08 |
0.74 |
0.85 |
| 16 |
A |
1357 |
1357 |
8.09 |
20.79 |
0.72 |
0.84 |
| 17 |
A |
1295 |
1295 |
4.77 |
9.00 |
0.74 |
0.85 |
| 18 |
A |
1265 |
1265 |
31.15 |
12.80 |
0.74 |
0.85 |
| 19 |
A |
1182 |
1182 |
6.10 |
3.49 |
0.61 |
0.76 |
| 20 |
A |
1137 |
1137 |
1.12 |
3.30 |
0.63 |
0.77 |
| 21 |
A |
1085 |
1085 |
32.91 |
4.16 |
0.65 |
0.79 |
| 22 |
A |
975 |
975 |
45.97 |
2.20 |
0.64 |
0.78 |
| 23 |
A |
952 |
952 |
28.32 |
4.20 |
0.71 |
0.83 |
| 24 |
A |
882 |
882 |
0.75 |
11.39 |
0.17 |
0.29 |
| 25 |
A |
799 |
799 |
2.02 |
1.40 |
0.34 |
0.51 |
| 26 |
A |
484 |
484 |
10.01 |
0.37 |
0.58 |
0.73 |
| 27 |
A |
333 |
333 |
12.34 |
0.67 |
0.60 |
0.75 |
| 28 |
A |
265 |
265 |
153.32 |
7.80 |
0.74 |
0.85 |
| 29 |
A |
229 |
229 |
4.24 |
0.13 |
0.72 |
0.84 |
| 30 |
A |
147 |
147 |
12.05 |
0.19 |
0.75 |
0.85 |
Unscaled Zero Point Vibrational Energy (zpe) 24148.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 24148.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.535 |
-0.530 |
0.134 |
| C2 |
-0.642 |
0.644 |
-0.298 |
| C3 |
0.760 |
0.576 |
0.297 |
| O4 |
1.387 |
-0.645 |
-0.216 |
| H5 |
-2.524 |
-0.463 |
-0.331 |
| H6 |
-1.674 |
-0.536 |
1.222 |
| H7 |
-1.076 |
-1.478 |
-0.156 |
| H8 |
-0.549 |
0.661 |
-1.391 |
| H9 |
-1.096 |
1.599 |
0.001 |
| H10 |
0.707 |
0.549 |
1.395 |
| H11 |
1.346 |
1.458 |
0.003 |
| H12 |
2.270 |
-0.771 |
0.183 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.5369 | 2.5528 | 2.9453 | 1.0952 | 1.0971 | 1.0928 | 2.1713 | 2.1775 | 2.7889 | 3.5023 | 3.8125 |
C2 | 1.5369 | | 1.5252 | 2.4060 | 2.1839 | 2.1839 | 2.1705 | 1.0966 | 1.0986 | 2.1672 | 2.1691 | 3.2731 | C3 | 2.5528 | 1.5252 | | 1.4649 | 3.5017 | 2.8315 | 2.7914 | 2.1376 | 2.1402 | 1.1001 | 1.0988 | 2.0258 | O4 | 2.9453 | 2.4060 | 1.4649 | | 3.9175 | 3.3836 | 2.6004 | 2.6144 | 3.3541 | 2.1174 | 2.1147 | 0.9761 | H5 | 1.0952 | 2.1839 | 3.5017 | 3.9175 | | 1.7724 | 1.7773 | 2.5077 | 2.5304 | 3.8003 | 4.3335 | 4.8311 | H6 | 1.0971 | 2.1839 | 2.8315 | 3.3836 | 1.7724 | | 1.7734 | 3.0864 | 2.5265 | 2.6216 | 3.8184 | 4.0848 | H7 | 1.0928 | 2.1705 | 2.7914 | 2.6004 | 1.7773 | 1.7734 | | 2.5248 | 3.0808 | 3.1128 | 3.8086 | 3.4357 | H8 | 2.1713 | 1.0966 | 2.1376 | 2.6144 | 2.5077 | 3.0864 | 2.5248 | | 1.7656 | 3.0579 | 2.4833 | 3.5311 | H9 | 2.1775 | 1.0986 | 2.1402 | 3.3541 | 2.5304 | 2.5265 | 3.0808 | 1.7656 | | 2.5091 | 2.4458 | 4.1202 | H10 | 2.7889 | 2.1672 | 1.1001 | 2.1174 | 3.8003 | 2.6216 | 3.1128 | 3.0579 | 2.5091 | | 1.7816 | 2.3779 | H11 | 3.5023 | 2.1691 | 1.0988 | 2.1147 | 4.3335 | 3.8184 | 3.8086 | 2.4833 | 2.4458 | 1.7816 | | 2.4192 | H12 | 3.8125 | 3.2731 | 2.0258 | 0.9761 | 4.8311 | 4.0848 | 3.4357 | 3.5311 | 4.1202 | 2.3779 | 2.4192 | |
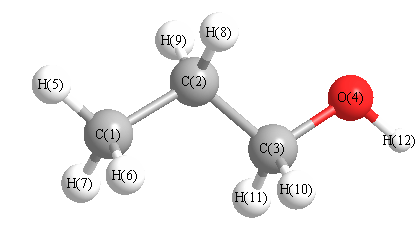 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.950 |
|
C1 |
C2 |
H8 |
109.946 |
| C1 |
C2 |
H9 |
110.318 |
|
C2 |
C1 |
H5 |
111.027 |
| C2 |
C1 |
H6 |
110.914 |
|
C2 |
C1 |
H7 |
110.109 |
| C2 |
C3 |
O4 |
107.135 |
|
C2 |
C3 |
H10 |
110.230 |
| C2 |
C3 |
H11 |
110.446 |
|
C3 |
C2 |
H8 |
108.125 |
| C3 |
C2 |
H9 |
108.210 |
|
C3 |
O4 |
H12 |
110.572 |
| O4 |
C3 |
H10 |
110.474 |
|
O4 |
C3 |
H11 |
110.328 |
| H5 |
C1 |
H6 |
107.897 |
|
H5 |
C1 |
H7 |
108.644 |
| H6 |
C1 |
H7 |
108.157 |
|
H8 |
C2 |
H9 |
107.093 |
| H10 |
C3 |
H11 |
108.237 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.407 |
|
|
|
| 2 |
C |
-0.248 |
|
|
|
| 3 |
C |
-0.015 |
|
|
|
| 4 |
O |
-0.636 |
|
|
|
| 5 |
H |
0.129 |
|
|
|
| 6 |
H |
0.123 |
|
|
|
| 7 |
H |
0.168 |
|
|
|
| 8 |
H |
0.149 |
|
|
|
| 9 |
H |
0.125 |
|
|
|
| 10 |
H |
0.122 |
|
|
|
| 11 |
H |
0.125 |
|
|
|
| 12 |
H |
0.364 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.833 |
1.134 |
1.216 |
1.859 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.554 |
-0.033 |
2.302 |
| y |
-0.033 |
-27.265 |
-0.723 |
| z |
2.302 |
-0.723 |
-27.051 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.604 |
-0.033 |
2.302 |
| y |
-0.033 |
-2.463 |
-0.723 |
| z |
2.302 |
-0.723 |
-2.142 |
|
| Polar |
|---|
| 3z2-r2 | -4.283 |
|---|
| x2-y2 | 4.711 |
|---|
| xy | -0.033 |
|---|
| xz | 2.302 |
|---|
| yz | -0.723 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.869 |
-0.137 |
0.140 |
| y |
-0.137 |
5.205 |
0.075 |
| z |
0.140 |
0.075 |
4.876 |
<r2> (average value of r
2) Å
2
| <r2> |
96.052 |
| (<r2>)1/2 |
9.801 |
