Jump to
S1C2
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -477.633375 |
| Energy at 298.15K | |
| HF Energy | -477.552196 |
| Nuclear repulsion energy | 105.066288 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3148 |
3148 |
27.86 |
|
|
|
| 2 |
A' |
3115 |
3115 |
16.41 |
|
|
|
| 3 |
A' |
3071 |
3071 |
21.10 |
|
|
|
| 4 |
A' |
2520 |
2520 |
55.33 |
|
|
|
| 5 |
A' |
1567 |
1567 |
4.54 |
|
|
|
| 6 |
A' |
1550 |
1550 |
3.72 |
|
|
|
| 7 |
A' |
1485 |
1485 |
6.39 |
|
|
|
| 8 |
A' |
1353 |
1353 |
49.93 |
|
|
|
| 9 |
A' |
1137 |
1137 |
2.08 |
|
|
|
| 10 |
A' |
1027 |
1027 |
6.44 |
|
|
|
| 11 |
A' |
834 |
834 |
5.46 |
|
|
|
| 12 |
A' |
625 |
625 |
3.60 |
|
|
|
| 13 |
A' |
303 |
303 |
4.40 |
|
|
|
| 14 |
A" |
3185 |
3185 |
29.54 |
|
|
|
| 15 |
A" |
3153 |
3153 |
5.80 |
|
|
|
| 16 |
A" |
1557 |
1557 |
9.07 |
|
|
|
| 17 |
A" |
1317 |
1317 |
0.59 |
|
|
|
| 18 |
A" |
1082 |
1082 |
0.06 |
|
|
|
| 19 |
A" |
823 |
823 |
5.41 |
|
|
|
| 20 |
A" |
251 |
251 |
0.81 |
|
|
|
| 21 |
A" |
153 |
153 |
27.90 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16627.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16627.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.524 |
0.752 |
0.000 |
| C2 |
0.000 |
0.870 |
0.000 |
| S3 |
-0.755 |
-0.887 |
0.000 |
| H4 |
1.974 |
1.751 |
0.000 |
| H5 |
1.879 |
0.218 |
0.887 |
| H6 |
1.879 |
0.218 |
-0.887 |
| H7 |
-0.358 |
1.389 |
0.891 |
| H8 |
-0.358 |
1.389 |
-0.891 |
| H9 |
-2.078 |
-0.504 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5284 | 2.8073 | 1.0955 | 1.0941 | 1.0941 | 2.1772 | 2.1772 | 3.8144 |
C2 | 1.5284 | | 1.9125 | 2.1614 | 2.1773 | 2.1773 | 1.0914 | 1.0914 | 2.4910 | S3 | 2.8073 | 1.9125 | | 3.7956 | 2.9910 | 2.9910 | 2.4767 | 2.4767 | 1.3770 | H4 | 1.0955 | 2.1614 | 3.7956 | | 1.7732 | 1.7732 | 2.5219 | 2.5219 | 4.6368 | H5 | 1.0941 | 2.1773 | 2.9910 | 1.7732 | | 1.7733 | 2.5248 | 3.0876 | 4.1186 | H6 | 1.0941 | 2.1773 | 2.9910 | 1.7732 | 1.7733 | | 3.0876 | 2.5248 | 4.1186 | H7 | 2.1772 | 1.0914 | 2.4767 | 2.5219 | 2.5248 | 3.0876 | | 1.7813 | 2.7086 | H8 | 2.1772 | 1.0914 | 2.4767 | 2.5219 | 3.0876 | 2.5248 | 1.7813 | | 2.7086 | H9 | 3.8144 | 2.4910 | 1.3770 | 4.6368 | 4.1186 | 4.1186 | 2.7086 | 2.7086 | |
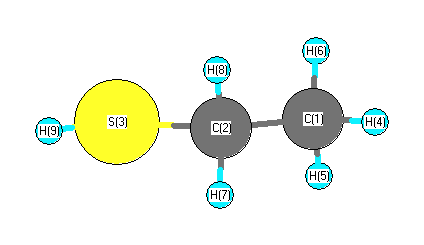 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
108.832 |
|
C1 |
C2 |
H7 |
111.323 |
| C1 |
C2 |
H8 |
111.323 |
|
C2 |
C1 |
H4 |
109.822 |
| C2 |
C1 |
H5 |
111.170 |
|
C2 |
C1 |
H6 |
111.170 |
| C2 |
S3 |
H9 |
97.101 |
|
S3 |
C2 |
H7 |
107.930 |
| S3 |
C2 |
H8 |
107.930 |
|
H4 |
C1 |
H5 |
108.154 |
| H4 |
C1 |
H6 |
108.154 |
|
H5 |
C1 |
H6 |
108.266 |
| H7 |
C2 |
H8 |
109.382 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.409 |
|
|
|
| 2 |
C |
-0.460 |
|
|
|
| 3 |
S |
0.008 |
|
|
|
| 4 |
H |
0.149 |
|
|
|
| 5 |
H |
0.160 |
|
|
|
| 6 |
H |
0.160 |
|
|
|
| 7 |
H |
0.175 |
|
|
|
| 8 |
H |
0.175 |
|
|
|
| 9 |
H |
0.041 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.126 |
2.092 |
0.000 |
2.096 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.810 |
-0.132 |
0.000 |
| y |
-0.132 |
-29.186 |
0.000 |
| z |
0.000 |
0.000 |
-29.328 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.447 |
-0.132 |
0.000 |
| y |
-0.132 |
-2.117 |
0.000 |
| z |
0.000 |
0.000 |
-2.330 |
|
| Polar |
|---|
| 3z2-r2 | -4.660 |
|---|
| x2-y2 | 4.376 |
|---|
| xy | -0.132 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.322 |
0.945 |
0.000 |
| y |
0.945 |
6.040 |
0.000 |
| z |
0.000 |
0.000 |
4.155 |
<r2> (average value of r
2) Å
2
| <r2> |
86.895 |
| (<r2>)1/2 |
9.322 |
Jump to
S1C1
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -477.634223 |
| Energy at 298.15K | -477.640505 |
| HF Energy | -477.552860 |
| Nuclear repulsion energy | 104.836091 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3184 |
3184 |
23.51 |
|
|
|
| 2 |
A |
3155 |
3155 |
9.99 |
|
|
|
| 3 |
A |
3135 |
3135 |
34.39 |
|
|
|
| 4 |
A |
3117 |
3117 |
8.67 |
|
|
|
| 5 |
A |
3062 |
3062 |
24.47 |
|
|
|
| 6 |
A |
2517 |
2517 |
50.92 |
|
|
|
| 7 |
A |
1563 |
1563 |
4.35 |
|
|
|
| 8 |
A |
1554 |
1554 |
10.74 |
|
|
|
| 9 |
A |
1543 |
1543 |
3.11 |
|
|
|
| 10 |
A |
1483 |
1483 |
5.03 |
|
|
|
| 11 |
A |
1352 |
1352 |
25.59 |
|
|
|
| 12 |
A |
1321 |
1321 |
2.47 |
|
|
|
| 13 |
A |
1146 |
1146 |
12.50 |
|
|
|
| 14 |
A |
1101 |
1101 |
0.27 |
|
|
|
| 15 |
A |
1019 |
1019 |
9.28 |
|
|
|
| 16 |
A |
874 |
874 |
13.11 |
|
|
|
| 17 |
A |
753 |
753 |
2.56 |
|
|
|
| 18 |
A |
617 |
617 |
7.45 |
|
|
|
| 19 |
A |
325 |
325 |
2.14 |
|
|
|
| 20 |
A |
259 |
259 |
1.42 |
|
|
|
| 21 |
A |
203 |
203 |
28.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16641.5 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16641.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.667 |
-0.366 |
-0.057 |
| C2 |
0.545 |
0.660 |
0.094 |
| S3 |
-1.200 |
-0.095 |
-0.080 |
| H4 |
2.642 |
0.132 |
0.005 |
| H5 |
1.625 |
-1.121 |
0.736 |
| H6 |
1.601 |
-0.880 |
-1.019 |
| H7 |
0.590 |
1.177 |
1.055 |
| H8 |
0.577 |
1.409 |
-0.700 |
| H9 |
-1.101 |
-0.965 |
0.983 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5275 | 2.8801 | 1.0968 | 1.0954 | 1.0935 | 2.1849 | 2.1794 | 3.0164 |
C2 | 1.5275 | | 1.9093 | 2.1644 | 2.1795 | 2.1743 | 1.0918 | 1.0921 | 2.4771 | S3 | 2.8801 | 1.9093 | | 3.8501 | 3.1153 | 3.0574 | 2.4722 | 2.4092 | 1.3775 | H4 | 1.0968 | 2.1644 | 3.8501 | | 1.7715 | 1.7770 | 2.5305 | 2.5277 | 4.0209 | H5 | 1.0954 | 2.1795 | 3.1153 | 1.7715 | | 1.7715 | 2.5405 | 3.0918 | 2.7418 | H6 | 1.0935 | 2.1743 | 3.0574 | 1.7770 | 1.7715 | | 3.0912 | 2.5279 | 3.3640 | H7 | 2.1849 | 1.0918 | 2.4722 | 2.5305 | 2.5405 | 3.0912 | | 1.7698 | 2.7296 | H8 | 2.1794 | 1.0921 | 2.4092 | 2.5277 | 3.0918 | 2.5279 | 1.7698 | | 3.3586 | H9 | 3.0164 | 2.4771 | 1.3775 | 4.0209 | 2.7418 | 3.3640 | 2.7296 | 3.3586 | |
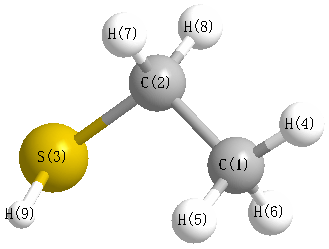 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
113.395 |
|
C1 |
C2 |
H7 |
111.985 |
| C1 |
C2 |
H8 |
111.517 |
|
C2 |
C1 |
H4 |
110.043 |
| C2 |
C1 |
H5 |
111.330 |
|
C2 |
C1 |
H6 |
111.027 |
| C2 |
S3 |
H9 |
96.473 |
|
S3 |
C2 |
H7 |
107.800 |
| S3 |
C2 |
H8 |
103.393 |
|
H4 |
C1 |
H5 |
107.822 |
| H4 |
C1 |
H6 |
108.448 |
|
H5 |
C1 |
H6 |
108.060 |
| H7 |
C2 |
H8 |
108.275 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.397 |
|
|
|
| 2 |
C |
-0.470 |
|
|
|
| 3 |
S |
0.011 |
|
|
|
| 4 |
H |
0.147 |
|
|
|
| 5 |
H |
0.143 |
|
|
|
| 6 |
H |
0.163 |
|
|
|
| 7 |
H |
0.177 |
|
|
|
| 8 |
H |
0.183 |
|
|
|
| 9 |
H |
0.043 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.939 |
0.012 |
0.967 |
2.167 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.923 |
1.452 |
-1.143 |
| y |
1.452 |
-27.354 |
-1.630 |
| z |
-1.143 |
-1.630 |
-27.534 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.478 |
1.452 |
-1.143 |
| y |
1.452 |
1.374 |
-1.630 |
| z |
-1.143 |
-1.630 |
1.104 |
|
| Polar |
|---|
| 3z2-r2 | 2.209 |
|---|
| x2-y2 | -2.568 |
|---|
| xy | 1.452 |
|---|
| xz | -1.143 |
|---|
| yz | -1.630 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.784 |
0.640 |
0.071 |
| y |
0.640 |
4.970 |
-0.522 |
| z |
0.071 |
-0.522 |
4.811 |
<r2> (average value of r
2) Å
2
| <r2> |
87.474 |
| (<r2>)1/2 |
9.353 |
