Jump to
S1C2
Energy calculated at wB97X-D/6-31G
| | hartrees |
|---|
| Energy at 0K | -477.919638 |
| Energy at 298.15K | |
| HF Energy | -477.919638 |
| Nuclear repulsion energy | 105.604270 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at wB97X-D/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3162 |
3162 |
25.20 |
|
|
|
| 2 |
A' |
3120 |
3120 |
18.01 |
|
|
|
| 3 |
A' |
3076 |
3076 |
21.78 |
|
|
|
| 4 |
A' |
2551 |
2551 |
50.70 |
|
|
|
| 5 |
A' |
1556 |
1556 |
6.66 |
|
|
|
| 6 |
A' |
1542 |
1542 |
4.70 |
|
|
|
| 7 |
A' |
1478 |
1478 |
11.03 |
|
|
|
| 8 |
A' |
1348 |
1348 |
43.57 |
|
|
|
| 9 |
A' |
1139 |
1139 |
2.62 |
|
|
|
| 10 |
A' |
1035 |
1035 |
8.00 |
|
|
|
| 11 |
A' |
838 |
838 |
6.32 |
|
|
|
| 12 |
A' |
652 |
652 |
3.67 |
|
|
|
| 13 |
A' |
308 |
308 |
5.02 |
|
|
|
| 14 |
A" |
3193 |
3193 |
27.46 |
|
|
|
| 15 |
A" |
3162 |
3162 |
5.12 |
|
|
|
| 16 |
A" |
1543 |
1543 |
11.98 |
|
|
|
| 17 |
A" |
1307 |
1307 |
0.78 |
|
|
|
| 18 |
A" |
1083 |
1083 |
0.00 |
|
|
|
| 19 |
A" |
827 |
827 |
6.67 |
|
|
|
| 20 |
A" |
236 |
236 |
0.27 |
|
|
|
| 21 |
A" |
17 |
17 |
32.68 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16585.2 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16585.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at wB97X-D/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.519 |
0.741 |
0.000 |
| C2 |
0.000 |
0.861 |
0.000 |
| S3 |
-0.753 |
-0.877 |
0.000 |
| H4 |
1.970 |
1.739 |
0.000 |
| H5 |
1.874 |
0.207 |
0.887 |
| H6 |
1.874 |
0.207 |
-0.887 |
| H7 |
-0.354 |
1.384 |
0.890 |
| H8 |
-0.354 |
1.384 |
-0.890 |
| H9 |
-2.071 |
-0.494 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5241 | 2.7902 | 1.0950 | 1.0940 | 1.0940 | 2.1713 | 2.1713 | 3.7966 |
C2 | 1.5241 | | 1.8942 | 2.1571 | 2.1734 | 2.1734 | 1.0917 | 1.0917 | 2.4744 | S3 | 2.7902 | 1.8942 | | 3.7767 | 2.9773 | 2.9773 | 2.4626 | 2.4626 | 1.3718 | H4 | 1.0950 | 2.1571 | 3.7767 | | 1.7726 | 1.7726 | 2.5139 | 2.5139 | 4.6168 | H5 | 1.0940 | 2.1734 | 2.9773 | 1.7726 | | 1.7736 | 2.5190 | 3.0829 | 4.1031 | H6 | 1.0940 | 2.1734 | 2.9773 | 1.7726 | 1.7736 | | 3.0829 | 2.5190 | 4.1031 | H7 | 2.1713 | 1.0917 | 2.4626 | 2.5139 | 2.5190 | 3.0829 | | 1.7810 | 2.6957 | H8 | 2.1713 | 1.0917 | 2.4626 | 2.5139 | 3.0829 | 2.5190 | 1.7810 | | 2.6957 | H9 | 3.7966 | 2.4744 | 1.3718 | 4.6168 | 4.1031 | 4.1031 | 2.6957 | 2.6957 | |
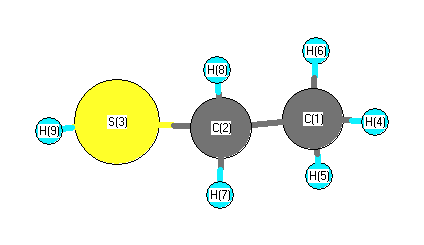 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
108.949 |
|
C1 |
C2 |
H7 |
111.140 |
| C1 |
C2 |
H8 |
111.140 |
|
C2 |
C1 |
H4 |
109.811 |
| C2 |
C1 |
H5 |
111.162 |
|
C2 |
C1 |
H6 |
111.162 |
| C2 |
S3 |
H9 |
97.221 |
|
S3 |
C2 |
H7 |
108.101 |
| S3 |
C2 |
H8 |
108.101 |
|
H4 |
C1 |
H5 |
108.147 |
| H4 |
C1 |
H6 |
108.147 |
|
H5 |
C1 |
H6 |
108.308 |
| H7 |
C2 |
H8 |
109.310 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.465 |
|
|
|
| 2 |
C |
-0.519 |
|
|
|
| 3 |
S |
0.003 |
|
|
|
| 4 |
H |
0.170 |
|
|
|
| 5 |
H |
0.181 |
|
|
|
| 6 |
H |
0.181 |
|
|
|
| 7 |
H |
0.198 |
|
|
|
| 8 |
H |
0.198 |
|
|
|
| 9 |
H |
0.053 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.157 |
2.177 |
0.000 |
2.182 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.183 |
-0.134 |
0.000 |
| y |
-0.134 |
-28.827 |
0.000 |
| z |
0.000 |
0.000 |
-28.884 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.673 |
-0.134 |
0.000 |
| y |
-0.134 |
-2.293 |
0.000 |
| z |
0.000 |
0.000 |
-2.379 |
|
| Polar |
|---|
| 3z2-r2 | -4.759 |
|---|
| x2-y2 | 4.644 |
|---|
| xy | -0.134 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.298 |
0.905 |
0.000 |
| y |
0.905 |
5.987 |
0.000 |
| z |
0.000 |
0.000 |
4.188 |
<r2> (average value of r
2) Å
2
| <r2> |
85.822 |
| (<r2>)1/2 |
9.264 |
Jump to
S1C1
Energy calculated at wB97X-D/6-31G
| | hartrees |
|---|
| Energy at 0K | -477.920670 |
| Energy at 298.15K | |
| HF Energy | -477.920670 |
| Nuclear repulsion energy | 105.423241 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at wB97X-D/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3189 |
3189 |
20.40 |
27.61 |
0.74 |
0.85 |
| 2 |
A |
3160 |
3160 |
10.78 |
88.06 |
0.74 |
0.85 |
| 3 |
A |
3142 |
3142 |
29.04 |
88.79 |
0.58 |
0.73 |
| 4 |
A |
3118 |
3118 |
11.37 |
109.23 |
0.18 |
0.31 |
| 5 |
A |
3062 |
3062 |
24.12 |
122.11 |
0.04 |
0.07 |
| 6 |
A |
2546 |
2546 |
46.81 |
140.91 |
0.38 |
0.55 |
| 7 |
A |
1550 |
1550 |
6.01 |
15.63 |
0.73 |
0.85 |
| 8 |
A |
1544 |
1544 |
14.24 |
22.09 |
0.75 |
0.86 |
| 9 |
A |
1534 |
1534 |
5.20 |
19.72 |
0.75 |
0.86 |
| 10 |
A |
1474 |
1474 |
9.42 |
5.76 |
0.68 |
0.81 |
| 11 |
A |
1350 |
1350 |
20.43 |
3.86 |
0.62 |
0.77 |
| 12 |
A |
1313 |
1313 |
2.45 |
11.59 |
0.75 |
0.86 |
| 13 |
A |
1146 |
1146 |
12.03 |
13.36 |
0.75 |
0.86 |
| 14 |
A |
1104 |
1104 |
0.45 |
7.34 |
0.55 |
0.71 |
| 15 |
A |
1028 |
1028 |
11.06 |
6.61 |
0.75 |
0.86 |
| 16 |
A |
881 |
881 |
15.39 |
8.73 |
0.75 |
0.86 |
| 17 |
A |
761 |
761 |
3.14 |
5.71 |
0.64 |
0.78 |
| 18 |
A |
645 |
645 |
7.77 |
24.47 |
0.31 |
0.47 |
| 19 |
A |
334 |
334 |
2.16 |
2.25 |
0.47 |
0.64 |
| 20 |
A |
257 |
257 |
2.16 |
0.55 |
0.70 |
0.82 |
| 21 |
A |
205 |
205 |
32.00 |
12.15 |
0.75 |
0.85 |
Unscaled Zero Point Vibrational Energy (zpe) 16671.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16671.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at wB97X-D/6-31G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.655 |
-0.364 |
-0.057 |
| C2 |
0.535 |
0.658 |
0.094 |
| S3 |
-1.191 |
-0.094 |
-0.079 |
| H4 |
2.629 |
0.135 |
0.007 |
| H5 |
1.612 |
-1.120 |
0.734 |
| H6 |
1.591 |
-0.876 |
-1.021 |
| H7 |
0.584 |
1.174 |
1.055 |
| H8 |
0.572 |
1.409 |
-0.697 |
| H9 |
-1.079 |
-0.983 |
0.962 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5233 | 2.8583 | 1.0961 | 1.0954 | 1.0935 | 2.1792 | 2.1738 | 2.9822 |
C2 | 1.5233 | | 1.8905 | 2.1601 | 2.1753 | 2.1703 | 1.0920 | 1.0921 | 2.4595 | S3 | 2.8583 | 1.8905 | | 3.8275 | 3.0936 | 3.0386 | 2.4584 | 2.3981 | 1.3729 | H4 | 1.0961 | 2.1601 | 3.8275 | | 1.7710 | 1.7768 | 2.5222 | 2.5203 | 3.9884 | H5 | 1.0954 | 2.1753 | 3.0936 | 1.7710 | | 1.7720 | 2.5348 | 3.0870 | 2.7042 | H6 | 1.0935 | 2.1703 | 3.0386 | 1.7768 | 1.7720 | | 3.0865 | 2.5226 | 3.3268 | H7 | 2.1792 | 1.0920 | 2.4584 | 2.5222 | 2.5348 | 3.0865 | | 1.7685 | 2.7249 | H8 | 2.1738 | 1.0921 | 2.3981 | 2.5203 | 3.0870 | 2.5226 | 1.7685 | | 3.3467 | H9 | 2.9822 | 2.4595 | 1.3729 | 3.9884 | 2.7042 | 3.3268 | 2.7249 | 3.3467 | |
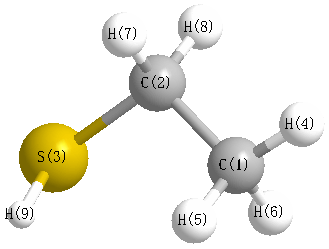 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
113.271 |
|
C1 |
C2 |
H7 |
111.810 |
| C1 |
C2 |
H8 |
111.373 |
|
C2 |
C1 |
H4 |
110.036 |
| C2 |
C1 |
H5 |
111.288 |
|
C2 |
C1 |
H6 |
111.004 |
| C2 |
S3 |
H9 |
96.530 |
|
S3 |
C2 |
H7 |
108.024 |
| S3 |
C2 |
H8 |
103.793 |
|
H4 |
C1 |
H5 |
107.826 |
| H4 |
C1 |
H6 |
108.477 |
|
H5 |
C1 |
H6 |
108.101 |
| H7 |
C2 |
H8 |
108.141 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.454 |
|
|
|
| 2 |
C |
-0.530 |
|
|
|
| 3 |
S |
0.007 |
|
|
|
| 4 |
H |
0.168 |
|
|
|
| 5 |
H |
0.162 |
|
|
|
| 6 |
H |
0.183 |
|
|
|
| 7 |
H |
0.200 |
|
|
|
| 8 |
H |
0.208 |
|
|
|
| 9 |
H |
0.054 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.032 |
-0.017 |
1.007 |
2.268 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.565 |
1.497 |
-1.182 |
| y |
1.497 |
-26.771 |
-1.673 |
| z |
-1.182 |
-1.673 |
-27.132 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.614 |
1.497 |
-1.182 |
| y |
1.497 |
1.578 |
-1.673 |
| z |
-1.182 |
-1.673 |
1.036 |
|
| Polar |
|---|
| 3z2-r2 | 2.072 |
|---|
| x2-y2 | -2.794 |
|---|
| xy | 1.497 |
|---|
| xz | -1.182 |
|---|
| yz | -1.673 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.669 |
0.608 |
0.078 |
| y |
0.608 |
5.000 |
-0.512 |
| z |
0.078 |
-0.512 |
4.801 |
<r2> (average value of r
2) Å
2
| <r2> |
86.264 |
| (<r2>)1/2 |
9.288 |
