return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C2H4N2S2 (Ethanedithioamide)

using model chemistry: MP2=FULL/6-31G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1AG
Energy calculated at MP2=FULL/6-31G
 hartrees
Energy at 0K-982.388631
Energy at 298.15K-982.394908
HF Energy-981.859117
Nuclear repulsion energy330.123582
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3697 3513 0.00      
2 Ag 3518 3342 0.00      
3 Ag 1706 1621 0.00      
4 Ag 1472 1399 0.00      
5 Ag 1370 1302 0.00      
6 Ag 977 928 0.00      
7 Ag 673 640 0.00      
8 Ag 434 412 0.00      
9 Ag 342 325 0.00      
10 Au 707 671 610.10      
11 Au 628 597 4.45      
12 Au 391 372 24.33      
13 Au 42 40 12.88      
14 Bg 728 691 0.00      
15 Bg 671 638 0.00      
16 Bg 628 597 0.00      
17 Bu 3699 3514 159.40      
18 Bu 3523 3347 249.25      
19 Bu 1689 1604 377.34      
20 Bu 1444 1372 383.33      
21 Bu 1283 1219 106.04      
22 Bu 851 809 57.67      
23 Bu 472 448 1.12      
24 Bu 283 269 21.81      

Unscaled Zero Point Vibrational Energy (zpe) 15613.5 cm-1
Scaled (by 0.9501) Zero Point Vibrational Energy (zpe) 14834.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-31G
ABC
0.14572 0.05078 0.03766

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-31G

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.059 0.757 0.000
C2 0.059 -0.757 0.000
S3 1.324 1.796 0.000
S4 -1.324 -1.796 0.000
N5 -1.324 1.229 0.000
N6 1.324 -1.229 0.000
H7 -2.107 0.579 0.000
H8 -1.485 2.228 0.000
H9 2.107 -0.579 0.000
H10 1.485 -2.228 0.000

Atom - Atom Distances (Å)
  C1 C2 S3 S4 N5 N6 H7 H8 H9 H10
C11.51941.72922.84971.35012.42002.05612.04882.54473.3612
C21.51942.84971.72922.42001.35012.54473.36122.05612.0488
S31.72922.84974.46242.70763.02503.64032.84152.50074.0278
S42.84971.72924.46243.02502.70762.50074.02783.64032.8415
N51.35012.42002.70763.02503.61251.01781.01243.87794.4543
N62.42001.35013.02502.70763.61253.87794.45431.01781.0124
H72.05612.54473.64032.50071.01783.87791.76314.37014.5586
H82.04883.36122.84154.02781.01244.45431.76314.55865.3555
H92.54472.05612.50073.64033.87791.01784.37014.55861.7631
H103.36122.04884.02782.84154.45431.01244.55865.35551.7631

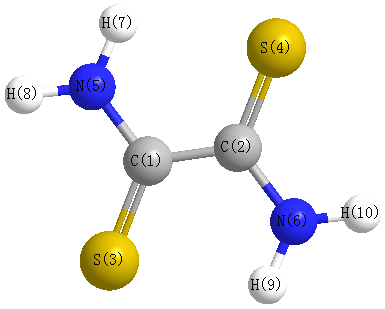
picture of Ethanedithioamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 S4 122.490 C1 C2 N6 114.869
C1 N5 H7 119.867 C1 N5 H8 119.586
C2 C1 S3 122.490 C2 C1 N5 114.869
C2 N6 H9 119.867 C2 N6 H10 119.586
S3 C1 N5 122.641 S4 C2 N6 122.641
H7 N5 H8 120.547 H9 N6 H10 120.547
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability