Jump to
S1C2
S1C3
Energy calculated at MP2=FULL/6-31G
| | hartrees |
|---|
| Energy at 0K | -169.140474 |
| Energy at 298.15K | -169.144400 |
| HF Energy | -168.806690 |
| Nuclear repulsion energy | 69.179042 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3669 |
3486 |
41.07 |
|
|
|
| 2 |
A' |
3468 |
3295 |
4.41 |
|
|
|
| 3 |
A' |
3087 |
2933 |
57.91 |
|
|
|
| 4 |
A' |
1718 |
1633 |
105.60 |
|
|
|
| 5 |
A' |
1439 |
1368 |
9.19 |
|
|
|
| 6 |
A' |
1298 |
1234 |
130.41 |
|
|
|
| 7 |
A' |
1172 |
1114 |
62.81 |
|
|
|
| 8 |
A' |
998 |
949 |
272.20 |
|
|
|
| 9 |
A' |
608 |
577 |
2.35 |
|
|
|
| 10 |
A" |
1048 |
996 |
6.77 |
|
|
|
| 11 |
A" |
842 |
800 |
81.77 |
|
|
|
| 12 |
A" |
355 |
337 |
112.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9850.8 cm
-1
Scaled (by 0.9501) Zero Point Vibrational Energy (zpe) 9359.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.401 |
0.000 |
| O2 |
-1.023 |
-0.553 |
0.000 |
| N3 |
1.220 |
-0.008 |
0.000 |
| H4 |
-0.351 |
1.442 |
0.000 |
| H5 |
-1.906 |
-0.131 |
0.000 |
| H6 |
1.900 |
0.763 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3991 | 1.2869 | 1.0979 | 1.9786 | 1.9341 |
O2 | 1.3991 | | 2.3086 | 2.1050 | 0.9784 | 3.2056 | N3 | 1.2869 | 2.3086 | | 2.1375 | 3.1283 | 1.0276 | H4 | 1.0979 | 2.1050 | 2.1375 | | 2.2115 | 2.3510 | H5 | 1.9786 | 0.9784 | 3.1283 | 2.2115 | | 3.9093 | H6 | 1.9341 | 3.2056 | 1.0276 | 2.3510 | 3.9093 | |
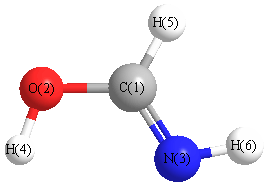 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
111.422 |
|
C1 |
N3 |
H6 |
112.887 |
| O2 |
C1 |
N3 |
118.452 |
|
O2 |
C1 |
H4 |
114.380 |
| N3 |
C1 |
H4 |
127.168 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at MP2=FULL/6-31G
| | hartrees |
|---|
| Energy at 0K | -169.153190 |
| Energy at 298.15K | -169.157322 |
| HF Energy | -168.820649 |
| Nuclear repulsion energy | 69.508026 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3585 |
3406 |
34.13 |
|
|
|
| 2 |
A' |
3494 |
3319 |
4.26 |
|
|
|
| 3 |
A' |
3198 |
3039 |
27.86 |
|
|
|
| 4 |
A' |
1673 |
1589 |
148.74 |
|
|
|
| 5 |
A' |
1409 |
1339 |
21.14 |
|
|
|
| 6 |
A' |
1385 |
1316 |
30.53 |
|
|
|
| 7 |
A' |
1170 |
1111 |
78.39 |
|
|
|
| 8 |
A' |
1016 |
965 |
231.19 |
|
|
|
| 9 |
A' |
572 |
544 |
53.71 |
|
|
|
| 10 |
A" |
1060 |
1007 |
0.51 |
|
|
|
| 11 |
A" |
844 |
802 |
67.35 |
|
|
|
| 12 |
A" |
633 |
602 |
257.88 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10019.1 cm
-1
Scaled (by 0.9501) Zero Point Vibrational Energy (zpe) 9519.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.441 |
0.000 |
| O2 |
-1.133 |
-0.362 |
0.000 |
| N3 |
1.182 |
-0.077 |
0.000 |
| H4 |
-0.295 |
1.491 |
0.000 |
| H5 |
-0.858 |
-1.309 |
0.000 |
| H6 |
1.946 |
0.607 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3894 | 1.2907 | 1.0900 | 1.9492 | 1.9530 |
O2 | 1.3894 | | 2.3328 | 2.0339 | 0.9857 | 3.2283 | N3 | 1.2907 | 2.3328 | | 2.1538 | 2.3831 | 1.0255 | H4 | 1.0900 | 2.0339 | 2.1538 | | 2.8555 | 2.4088 | H5 | 1.9492 | 0.9857 | 2.3831 | 2.8555 | | 3.3960 | H6 | 1.9530 | 3.2283 | 1.0255 | 2.4088 | 3.3960 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
109.126 |
|
C1 |
N3 |
H6 |
114.478 |
| O2 |
C1 |
N3 |
120.977 |
|
O2 |
C1 |
H4 |
109.644 |
| N3 |
C1 |
H4 |
129.379 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at MP2=FULL/6-31G
| | hartrees |
|---|
| Energy at 0K | -169.147859 |
| Energy at 298.15K | -169.151880 |
| HF Energy | -168.813240 |
| Nuclear repulsion energy | 68.780015 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3600 |
3421 |
16.91 |
|
|
|
| 2 |
A' |
3402 |
3232 |
10.77 |
|
|
|
| 3 |
A' |
3286 |
3122 |
8.93 |
|
|
|
| 4 |
A' |
1647 |
1565 |
133.12 |
|
|
|
| 5 |
A' |
1436 |
1364 |
0.03 |
|
|
|
| 6 |
A' |
1314 |
1248 |
17.02 |
|
|
|
| 7 |
A' |
1114 |
1058 |
254.03 |
|
|
|
| 8 |
A' |
995 |
945 |
146.18 |
|
|
|
| 9 |
A' |
561 |
533 |
35.27 |
|
|
|
| 10 |
A" |
1096 |
1041 |
103.18 |
|
|
|
| 11 |
A" |
835 |
793 |
51.55 |
|
|
|
| 12 |
A" |
492 |
467 |
145.82 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9888.1 cm
-1
Scaled (by 0.9501) Zero Point Vibrational Energy (zpe) 9394.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.452 |
0.000 |
| O2 |
-1.133 |
-0.388 |
0.000 |
| N3 |
1.257 |
0.158 |
0.000 |
| H4 |
-0.330 |
1.484 |
0.000 |
| H5 |
-0.892 |
-1.342 |
0.000 |
| H6 |
1.488 |
-0.849 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.4105 | 1.2909 | 1.0838 | 2.0036 | 1.9761 |
O2 | 1.4105 | | 2.4514 | 2.0374 | 0.9836 | 2.6607 | N3 | 1.2909 | 2.4514 | | 2.0683 | 2.6208 | 1.0327 | H4 | 1.0838 | 2.0374 | 2.0683 | | 2.8817 | 2.9575 | H5 | 2.0036 | 0.9836 | 2.6208 | 2.8817 | | 2.4305 | H6 | 1.9761 | 2.6607 | 1.0327 | 2.9575 | 2.4305 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
112.402 |
|
C1 |
N3 |
H6 |
116.079 |
| O2 |
C1 |
N3 |
130.269 |
|
O2 |
C1 |
H4 |
108.832 |
| N3 |
C1 |
H4 |
120.899 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
