Jump to
S1C2
S1C3
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -169.573347 |
| Energy at 298.15K | -169.577298 |
| HF Energy | -169.460589 |
| Nuclear repulsion energy | 69.682133 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3686 |
3686 |
35.63 |
|
|
|
| 2 |
A' |
3497 |
3497 |
4.22 |
|
|
|
| 3 |
A' |
3084 |
3084 |
63.05 |
|
|
|
| 4 |
A' |
1751 |
1751 |
139.68 |
|
|
|
| 5 |
A' |
1441 |
1441 |
11.18 |
|
|
|
| 6 |
A' |
1293 |
1293 |
140.88 |
|
|
|
| 7 |
A' |
1174 |
1174 |
52.45 |
|
|
|
| 8 |
A' |
1000 |
1000 |
286.88 |
|
|
|
| 9 |
A' |
615 |
615 |
2.55 |
|
|
|
| 10 |
A" |
1059 |
1059 |
5.56 |
|
|
|
| 11 |
A" |
839 |
839 |
85.70 |
|
|
|
| 12 |
A" |
373 |
373 |
105.29 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9906.2 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 9906.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.395 |
0.000 |
| O2 |
-1.015 |
-0.550 |
0.000 |
| N3 |
1.211 |
-0.003 |
0.000 |
| H4 |
-0.353 |
1.431 |
0.000 |
| H5 |
-1.897 |
-0.133 |
0.000 |
| H6 |
1.894 |
0.758 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3872 | 1.2749 | 1.0949 | 1.9690 | 1.9286 |
O2 | 1.3872 | | 2.2927 | 2.0892 | 0.9753 | 3.1900 | N3 | 1.2749 | 2.2927 | | 2.1225 | 3.1108 | 1.0227 | H4 | 1.0949 | 2.0892 | 2.1225 | | 2.1979 | 2.3460 | H5 | 1.9690 | 0.9753 | 3.1108 | 2.1979 | | 3.8943 | H6 | 1.9286 | 3.1900 | 1.0227 | 2.3460 | 3.8943 | |
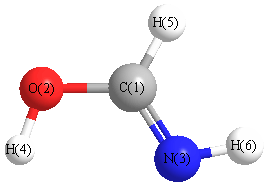 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
111.730 |
|
C1 |
N3 |
H6 |
113.695 |
| O2 |
C1 |
N3 |
118.850 |
|
O2 |
C1 |
H4 |
114.132 |
| N3 |
C1 |
H4 |
127.017 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.220 |
|
|
|
| 2 |
O |
-0.554 |
|
|
|
| 3 |
N |
-0.450 |
|
|
|
| 4 |
H |
0.132 |
|
|
|
| 5 |
H |
0.381 |
|
|
|
| 6 |
H |
0.270 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.867 |
3.790 |
0.000 |
4.225 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-13.184 |
1.452 |
0.000 |
| y |
1.452 |
-18.621 |
0.000 |
| z |
0.000 |
0.000 |
-18.764 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.508 |
1.452 |
0.000 |
| y |
1.452 |
-2.646 |
0.000 |
| z |
0.000 |
0.000 |
-2.862 |
|
| Polar |
|---|
| 3z2-r2 | -5.724 |
|---|
| x2-y2 | 5.436 |
|---|
| xy | 1.452 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.045 |
0.065 |
0.000 |
| y |
0.065 |
2.770 |
0.000 |
| z |
0.000 |
0.000 |
1.174 |
<r2> (average value of r
2) Å
2
| <r2> |
42.353 |
| (<r2>)1/2 |
6.508 |
Jump to
S1C1
S1C3
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -169.573347 |
| Energy at 298.15K | -169.577298 |
| HF Energy | -169.460589 |
| Nuclear repulsion energy | 69.682133 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G
Geometric Data calculated at B2PLYP/6-31G
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at B2PLYP/6-31G
| | hartrees |
|---|
| Energy at 0K | -169.580415 |
| Energy at 298.15K | -169.584458 |
| HF Energy | -169.466936 |
| Nuclear repulsion energy | 69.314472 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/6-31G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3618 |
3618 |
13.82 |
|
|
|
| 2 |
A' |
3425 |
3425 |
11.11 |
|
|
|
| 3 |
A' |
3282 |
3282 |
8.45 |
|
|
|
| 4 |
A' |
1684 |
1684 |
173.98 |
|
|
|
| 5 |
A' |
1434 |
1434 |
0.03 |
|
|
|
| 6 |
A' |
1308 |
1308 |
19.06 |
|
|
|
| 7 |
A' |
1102 |
1102 |
268.41 |
|
|
|
| 8 |
A' |
1013 |
1013 |
131.71 |
|
|
|
| 9 |
A' |
563 |
563 |
34.82 |
|
|
|
| 10 |
A" |
1092 |
1092 |
98.57 |
|
|
|
| 11 |
A" |
844 |
844 |
58.34 |
|
|
|
| 12 |
A" |
513 |
513 |
137.30 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9938.5 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 9938.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/6-31G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.448 |
0.000 |
| O2 |
-1.123 |
-0.383 |
0.000 |
| N3 |
1.246 |
0.155 |
0.000 |
| H4 |
-0.329 |
1.478 |
0.000 |
| H5 |
-0.891 |
-1.335 |
0.000 |
| H6 |
1.484 |
-0.845 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3968 | 1.2796 | 1.0814 | 1.9930 | 1.9685 |
O2 | 1.3968 | | 2.4289 | 2.0233 | 0.9803 | 2.6480 | N3 | 1.2796 | 2.4289 | | 2.0565 | 2.6045 | 1.0282 | H4 | 1.0814 | 2.0233 | 2.0565 | | 2.8687 | 2.9469 | H5 | 1.9930 | 0.9803 | 2.6045 | 2.8687 | | 2.4248 | H6 | 1.9685 | 2.6480 | 1.0282 | 2.9469 | 2.4248 | |
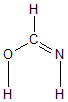 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
112.766 |
|
C1 |
N3 |
H6 |
116.650 |
| O2 |
C1 |
N3 |
130.284 |
|
O2 |
C1 |
H4 |
108.789 |
| N3 |
C1 |
H4 |
120.927 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/6-31G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.219 |
|
|
|
| 2 |
O |
-0.569 |
|
|
|
| 3 |
N |
-0.467 |
|
|
|
| 4 |
H |
0.190 |
|
|
|
| 5 |
H |
0.371 |
|
|
|
| 6 |
H |
0.256 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.495 |
-2.259 |
0.000 |
2.313 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.754 |
-2.378 |
0.000 |
| y |
-2.378 |
-13.060 |
0.000 |
| z |
0.000 |
0.000 |
-18.787 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.831 |
-2.378 |
0.000 |
| y |
-2.378 |
8.211 |
0.000 |
| z |
0.000 |
0.000 |
-0.380 |
|
| Polar |
|---|
| 3z2-r2 | -0.760 |
|---|
| x2-y2 | -10.695 |
|---|
| xy | -2.378 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.228 |
-0.102 |
0.000 |
| y |
-0.102 |
3.369 |
0.000 |
| z |
0.000 |
0.000 |
1.217 |
<r2> (average value of r
2) Å
2
| <r2> |
42.855 |
| (<r2>)1/2 |
6.546 |
